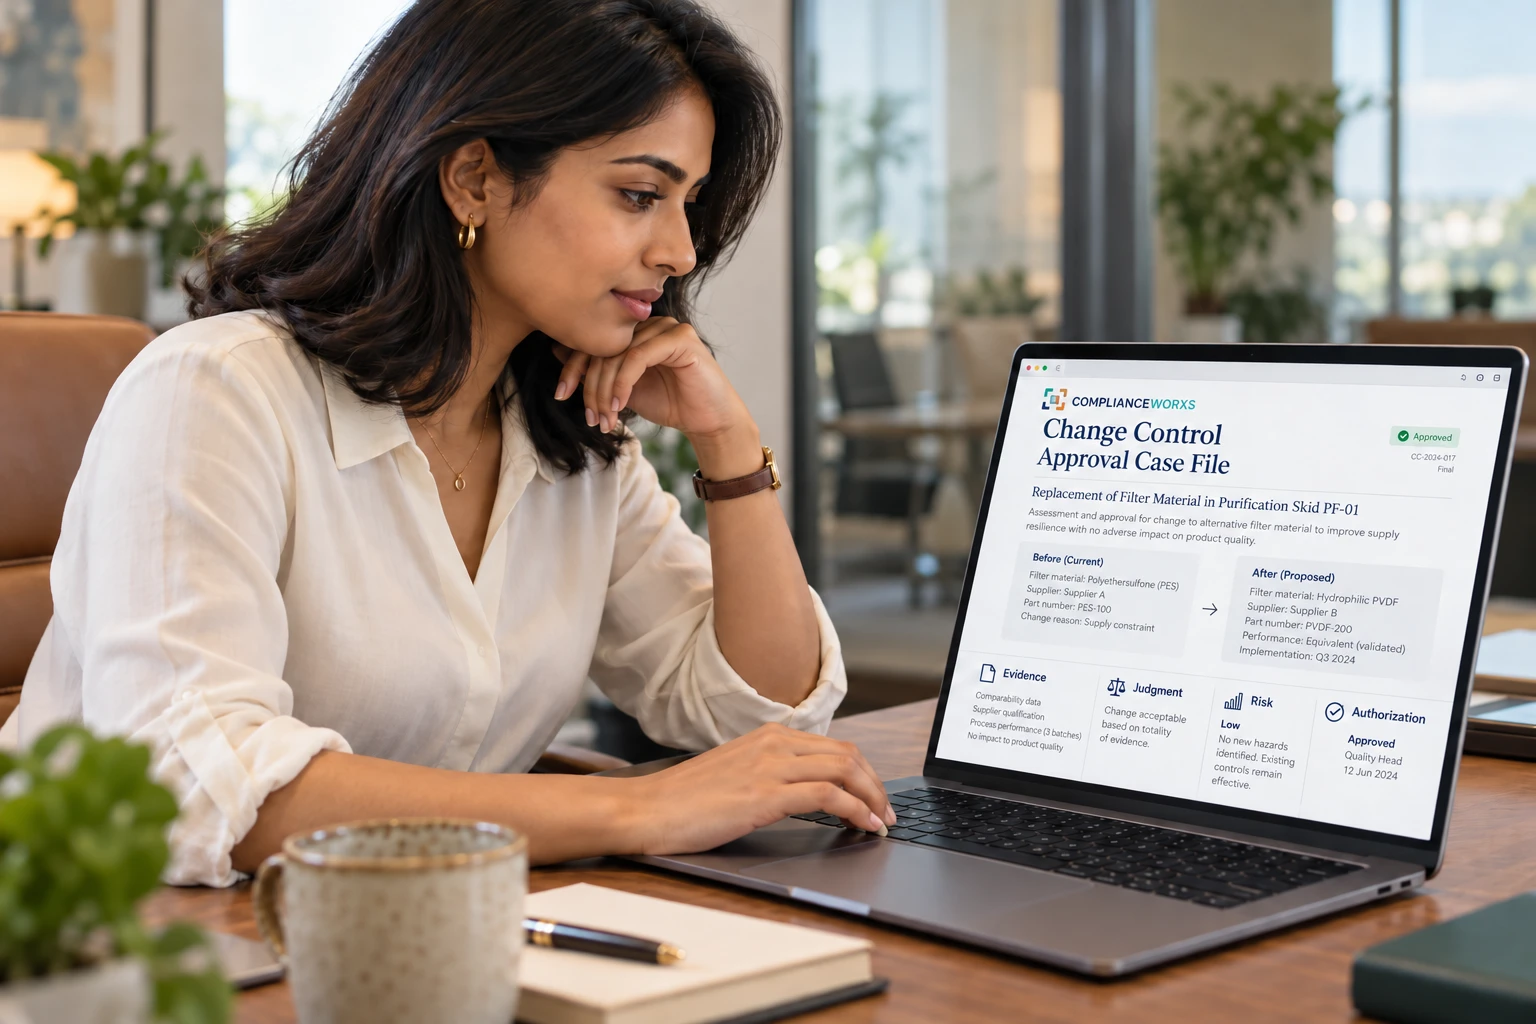
A risk category and a QA approval show that a change workflow was completed. The Case File shows what the decision needs to preserve: the evidence, the regulatory determination and its basis, the risk accepted, and who authorized implementation before it began.
When the inspector asks who authorized this change and on what basis, this is the record that answers.
Illustrative case. This is an excerpt of the worked record inside the Case File: Change Control CC-2025-0142, a proposed expansion of the granulation loss-on-drying range for an approved modified-release tablet, from 1.5%–2.5% to 1.5%–3.5%. The company, product, people, dates, identifiers, cost figures and events are fictional. This is not an FDA observation or a Form 483. In the scenario the firm implemented the change without this record. The excerpt shows the record that should have existed before implementation.
DecisionDo not implement or distribute; a Prior Approval Supplement is required first
Decision ownerElena Park, Vice President, Regulatory Affairs (illustrative)
Authorized12 June 2025, 14:40 EDT, before planned implementation
What was checked, and against which records
Approved application compared. The filed granulation LOD range is 1.5%–2.5%. The proposed upper limit of 3.5% falls outside it.
Development history. Moisture level is linked to compression behavior and modified-release performance.
Pilot data. Coverage at the proposed upper range is insufficient to establish dissolution comparability or continued state of control.
Quality-risk assessment. Dissolution, content uniformity, hardness and friability, stability, and process variability.
Regulatory assessment. No approved design space or comparability protocol covers the expansion.
The conclusion, and what was rejected
Major change: Prior Approval Supplement required. The change is treated as having substantial potential to affect product quality. Implementation is not authorized, and distribution of product made with the change is prohibited until FDA approval and site release conditions are complete.
Classify as Minor. Rejected: a selected category with no analysis behind it could not show why no filing was needed.
Treat as a tighter limit inside the filed range. Rejected: this extends the process beyond the application’s upper boundary.
A PAS is this scenario’s conclusion, not a rule for every range change. Reporting categories are fact-specific.
What must happen before it can proceed
Technical. A protocol-driven comparability package at the proposed range, with justified acceptance criteria for every relevant quality attribute.
Regulatory. Confirm the reporting category against the approved application, current regulations, FDA guidance and product commitments.
Quality. Approve revised procedures and batch records through the quality-unit controls.
System. Verify FDA approval status before the implementation hold is removed.
Post-change. Confirm the objective was met and no deleterious effect occurred.
Who authorized the decision
Elena Park, Vice President, Regulatory Affairs (illustrative), who classified the change and required the supplement.
System action. A regulatory hold was applied to the implementation task, the master batch record revision, and affected-material disposition.
The record shows who held the authority, what they relied on, and when they decided, before planned implementation. No one reconstructs it after the batches ship.
What the full Case File adds
The whole record, not one page of it.
One completed change-control decision record: context, evidence, judgment, risk and authority in one document.
The decision gap: what existed in the change file and what was missing.
The inspection consequence: distribution hold, a 36-month retrospective review, and an illustrative response cost above $2.4 million.
The authorization sequence, with the quality decision and the regulatory decision side by side.
The preventive controls that make the correct path unavoidable.
Rules and sources: 21 CFR 211.100(a), 21 CFR 314.70, FDA change guidance and ICH Q10 §3.2.3.
A PDF, delivered after payment, with a 7-day secure download.
A completed approval workflow is not the same decision as the authorization behind it.
A change can have a request, a risk category, an implementation plan, and a QA approval without preserving what evidence supported the change, which regulatory requirements governed it, or who authorized implementation. A classification field records an output. An authorization record preserves the basis: the facts considered, the quality-risk conclusion, the filing pathway, the implementation timing, the accountable owner, and the conditions that must be met before execution.
The two decisions
Change control needs a quality decision and a regulatory decision. Neither can be inferred from the other.
The product-quality decision
Which quality attributes and process controls could be affected?
What pre-change and proposed-state evidence is required?
Do the data show equivalence and continued state of control?
What acceptance criteria and post-change checks apply?
Who owns the scientific and quality conclusion?
The regulatory-filing decision
Is the change beyond the variations already provided for in the application?
What is its potential to adversely affect identity, strength, quality, purity or potency?
Which pathway applies: PAS, CBE-30, CBE-0 or annual report?
When may implementation and distribution lawfully occur?
Who owns the filing determination and the submission-status check?
The release rule. The system should release implementation only when the documented decision condition is satisfied, not merely when the approval workflow is complete. The test: could the system prove, without reconstruction, why this change was allowed to proceed on that date?
A risk category is marked, but the record does not show the analysis that supports it.
Filing impact not determined
The change was approved for implementation, but the record does not show that it was compared to the filed parameter ranges, or what filing it required.
Quality impact not shown
The record does not show which critical quality attributes were evaluated against the change, or the data behind that evaluation.
Approver not shown
The record carries a QA stamp, but not the named person who authorized the classification and the filing determination, or when they did so.
Your answers, and where the Case File answers them
Inside the $149 Case File
The change was implemented. Now explain the authorization.
See how one worked example connects the evidence, the reasoning, and accountable approval before implementation.
The setup · illustrative case
An investigator selects a change after commercial batches were made under an expanded granulation range, and asks who authorized implementation, what evidence showed the product would not be affected, and what basis supported the filing category before distribution. The record held a “Minor” category and a QA approval, and nothing behind either. In this scenario that means a distribution hold, a 36-month review of related changes, and a response cost above $2.4 million (an assumption of the case, not a verified figure). Here is what the record would have said.
In the inspector’s chair
The inspector asks
“What evidence showed the change would not adversely affect the product?”
From the record’s Evidence Considered
Approved application: granulation LOD operating range 1.5%–2.5%. Proposed master batch record: upper limit increased to 3.5%.
Development history linking moisture level to compression behavior and modified-release performance.
Available pilot data: insufficient coverage at the proposed upper range to establish dissolution comparability and continued state of control.
Why it matters. It separates the evidence that exists from the evidence still owed, and it shows the gap before any batch is made.
The inspector asks
“What specific basis supported the filing category before distribution?”
From the record’s Decision Basis
The proposed range expansion is not merely a tighter operating limit within the filed range. It extends the process beyond the application-described upper boundary for a modified-release product, while the available package does not adequately establish the absence of an adverse effect.
Under the facts assumed in this case, the change has substantial potential to affect product quality. A PAS is therefore required before distribution.
Why it matters. The filing category is tied to the filed range and the evidence, and it states the assumptions it depends on.
The inspector asks
“Who authorized implementation, and under what condition?”
From the record’s Decision Statement
Based on the evidence and assumptions listed, I classify the proposed change as a major change and require a Prior Approval Supplement. Implementation is not authorized. Distribution of product made using the change is prohibited until FDA approval and completion of site release conditions.
Elena Park · Vice President, Regulatory Affairs (illustrative) 12 June 2025 · 14:40 EDT · before planned implementation
Why it matters. A name, a title, a timestamp that precedes implementation, and a condition the system enforces. A QA approval shows the workflow finished; this is the authorization.
1 of 3
Get the Case File
Start with the finished approval record.
The Case File is a worked example for leaders and review teams. It does not document your own change controls. When an actual consequential decision needs its own record, the Inspection Response Record is built for that.